
Batch Manufacturing Record (BMR)
Batch Manufacturing Record requirements explained through master control, material genealogy, execution evidence, worked calculations and quality review.

On this page · 11 sections
- 1What is a Batch Manufacturing Record?
- 2What the pharmaceutical batch record must contain
- 3Master version control, issuance and batch identity
- 4Materials, weighing and genealogy
- 5Execution evidence: people, equipment and process conditions
- 6Three worked examples of batch record reasoning
- 7Data integrity in paper, electronic and hybrid records
- 8Deviation management, investigation and release review
- 9ERP, MES, WMS and laboratory integration
- 10Designing a usable BMR template and introducing it
- 11Retention, retrieval and meaningful performance measures
How does BMR apply to your shop floor?
Pick your industry and scale — Ask V5 rewrites the definition in your context, gives a worked example, and shows what V5 does on day one.
01What is a Batch Manufacturing Record?
A Batch Manufacturing Record (BMR) is the batch-specific history of manufacturing and control activities for a product. It connects the approved instructions with the materials, equipment, people, measurements, checks and decisions used for one identifiable batch. Its purpose is to make the manufacturing history reconstructable: a reviewer should be able to understand what was intended, what occurred, where execution differed and how the quality decision was reached.
The Master Manufacturing Record defines the approved process. The BMR records execution of that process. The Bill of Materials describes required inputs; it is one part of the manufacturing definition and does not replace operating instructions, sampling requirements or acceptance criteria.
A BMR can comprise linked records rather than one enormous document. Scale readings, equipment logs, laboratory results and investigation records may remain in their controlled systems, provided the batch record identifies them reliably and they remain available with the context needed for review. A printed summary containing only final values may be inadequate where original electronic data or metadata are necessary to understand a result.
“Batch record,” “batch production record” and “batch manufacturing record” often describe overlapping concepts, but terminology depends on sector and local procedures. BPR can also mean batch packaging record. Define the acronym in procedures and integrations; otherwise an interface can exchange the wrong document while appearing to use the correct name.
This guide concentrates on the operational BMR and US finished pharmaceutical requirements. Dietary supplements have their own batch-record provisions. Food, cosmetics, devices and other products require an applicability assessment rather than an assumption that every requirement in pharmaceutical GMP applies unchanged. The practical principles of identity, contemporaneous recording and traceable review are broadly useful; the legal basis must still be identified for the particular activity.
An electronic batch record changes how this evidence is captured and controlled. It does not change the need to demonstrate what happened. Paper, electronic and hybrid records should all make it possible to connect the batch to its master, execution data, exceptions and disposition.
Master record, BMR and genealogy: different questions
| Record | Question it answers | Evidence boundary |
|---|---|---|
| Master manufacturing record | What process and criteria are approved? | Controlled instructions, quantities and limits for the applicable version. |
| Batch Manufacturing Record | What happened for this batch? | Execution, results, exceptions, checks and disposition linked to that master. |
| Material genealogy | Which lots went into which outputs? | Input/output relationships and transformations; does not replace process or quality evidence. |
02What the pharmaceutical batch record must contain
For US finished pharmaceuticals, 21 CFR 211.188 requires batch production and control records for each batch. The record must include an accurate reproduction of the appropriate master production or control record, checked for accuracy, dated and signed. It must also document completion of significant manufacturing, processing, packing and holding steps.
The listed execution evidence includes dates, major equipment and lines, the identity of component and in-process material batches, quantities used, in-process and laboratory results, packaging and labeling area inspections, yields, labeling controls, containers and closures, sampling, identified personnel, investigations and relevant examinations of packaged products. A checklist can help map those requirements, but it should preserve the actual subsection references rather than recite an invented list.
The regulation's list does not say that every batch needs an independent handwritten narrative of every low-level machine event. Determine which evidence demonstrates significant steps and process control, then preserve it in a suitable form. A mixing operation may need its duration, speed, temperature profile and outcome; collecting thousands of irrelevant tags does not compensate for a missing start authorization or unidentified material lot.
21 CFR 211.186 addresses the master record. Its requirements include product identity, component information, quantities, theoretical yields and investigation limits, packaging information and manufacturing and control instructions. Read the master and batch requirements together: a batch record cannot establish conformity when the approved master omits the relevant criteria.
21 CFR 211.192 requires quality control unit review and approval of production and control records before release or distribution. It also requires thorough investigation of unexplained discrepancies and specification failures, with consideration of potentially associated batches and products. A clean final test does not erase an unresolved manufacturing discrepancy.
For dietary supplements, 21 CFR 111.260 specifies batch record contents. Link the product's regulatory classification to its record requirements instead of using a drug template with the headings renamed. Other markets may require additional documentation, release responsibilities or retention arrangements. Treat that mapping as part of controlled process design, with identifiable legal provisions and clearly separated company practices.
03Master version control, issuance and batch identity
The correct master is the starting point for a credible BMR. At issuance, identify the product, strength or configuration, intended batch size, master version and production order. Confirm that the selected version is approved and effective for the intended operation. Where several markets or presentations share a product code, the record also needs enough configuration information to distinguish the applicable formula, labeling and packaging instructions.
An electronic implementation commonly preserves a snapshot or immutable reference to the approved manufacturing definition. This is a design approach for controlling historical context, not a claim that the regulation prescribes one particular database architecture. The requirement is that reviewers can determine what instructions applied and examine an accurate, controlled representation of them.
Recipe versioning becomes particularly important when the master changes during a batch. Updating the current recipe must not silently rewrite already executed steps or alter what the historical batch appears to have required. Decide whether an approved change applies to future batches, an unstarted issued record or the remaining steps of a current batch. Preserve the authorization and impact assessment for the selected path.
A sound issuance process prevents accidental duplicate records and establishes how withdrawn, cancelled or superseded records are accounted for. If an order is cancelled after weighing has begun, the materials and intermediate containers still need a traceable disposition. Cancelling the administrative order is not proof that the physical material returned to inventory.
Use identifiers that survive movement across systems. The ERP production order, MES batch identifier, WMS handling-unit number and laboratory sample number may differ. Store explicit relationships between them rather than relying on matching descriptions. Reviewers should not have to guess that “Trial Blend 3” in a laboratory notebook is the same material as an unfamiliar ERP order.
Keep planned instructions and actual execution values distinct. The approved target, permitted tolerance and measured result answer different questions. A field that changes from target to actual after execution conceals the comparison a reviewer needs. Similarly, actual equipment should be recorded even where a preferred equipment class appears in the master.
Change control should cover changes to record templates as well as changes to formulas. A revised rounding rule, sampling field or signature step can affect the evidence generated even when the manufacturing recipe remains unchanged.
04Materials, weighing and genealogy
Material identity is where a BMR connects manufacturing to the supply chain. Record the actual component or intermediate lot, quantity and unit, together with the relevant operation. A material code alone cannot establish lot genealogy; a supplier name alone cannot establish which delivery was consumed. When a lot is split across containers, preserve the relationship between the lot and the handling units used.
Before consumption, verify the material's eligibility under the approved process: identity, quality status, expiry or retest conditions where relevant, and any product-specific restrictions. The BMR should preserve enough context to understand the eligibility decision at the time of use. The fact that a lot is released today does not establish that it was released when the operator selected it last month.
Weigh and dispense evidence should distinguish gross, tare and net measurements when those values are relevant to the operation. Record the unit and the scale identity. A value of “12.5” without a unit or attributable source is difficult to defend. Capture the actual result and acceptance decision rather than recording only a green pass indicator.
An electronic weighing record may also include the target, tolerance, stable-reading status and calculation version. Whether these belong in the batch summary or a linked source record depends on the process and the information needed to reconstruct the transaction. Preserve the source context when printing or exporting; a screen capture of the final number can omit the metadata that explains it.
Where strength corrections are required, distinguish the laboratory assay basis from the physical mass dispensed. The approved algorithm, input result, basis, unit conversions and calculated charge should be traceable. Do not let a free-text potency entry replace a controlled analytical result. A repeat calculation must use the authorized result and method, with a reason for any changed input.
Lot genealogy should reflect actual consumption, not merely allocations. Material reserved for an order can be returned, substituted or never used. Likewise, a partially consumed container may supply several batches. The record needs the quantities and event relationships that distinguish these cases.
Avoid overloading the BMR with supplier paperwork that is better managed elsewhere. A controlled reference to the approved component record and relevant Certificate of Analysis can be more useful than repeatedly attaching an unversioned copy. The reference must remain resolvable throughout retention, including when the supplier portal or file location changes.
05Execution evidence: people, equipment and process conditions
A manufacturing step should produce a clear record of its completion and outcome. Identify the person performing the work and any required direct supervision or check. For significant steps performed by automated equipment, the pharmaceutical rule addresses identification of the person checking the step. Do not reduce automated execution to an unexplained system username with no accountable review.
Design the record around the process sequence while accommodating legitimate parallel work. Material preparation, equipment readiness and laboratory sampling can occur on different timelines. A chronological event history combined with operation identifiers often makes these relationships clearer than a rigid list of pages.
Equipment evidence should identify the asset or line actually used and link to applicable readiness records. Line clearance, cleaning status, calibration and maintenance can affect eligibility, but the required checks depend on the operation. Record the decision and relevant evidence instead of copying a generic checklist into every step.
Process values need context. Temperature requires a unit and measurement point; pressure may be absolute or gauge; a mixing duration needs a defined start and finish event. An isolated maximum value may not establish compliance with a required time-temperature profile. When the process depends on trends, preserve the relevant series or a validated, traceable representation of it.
Sampling records should identify the sample, source batch, operation or location, timing and method reference as appropriate. The laboratory result must remain linked to the sample actually taken. A compliant result from a different sampling point cannot be substituted simply because the product and batch descriptions appear similar.
During execution, unexpected events should be recorded when they occur, with a reference to the relevant investigation or deviation process. A stop, alarm, transfer, manual adjustment or rejected reading may affect interpretation even if later values meet limits. Retain the chronology rather than editing the record into a smooth narrative after manufacturing ends.
Usability matters because cumbersome records encourage workarounds. Present the instructions and critical fields at the point of action. Avoid forcing operators to remember values for later transcription. Where a paper fallback is authorized, define its controlled issuance, recording and reconciliation with the electronic system. Training should cover the exceptional path as well as normal execution.
06Three worked examples of batch record reasoning
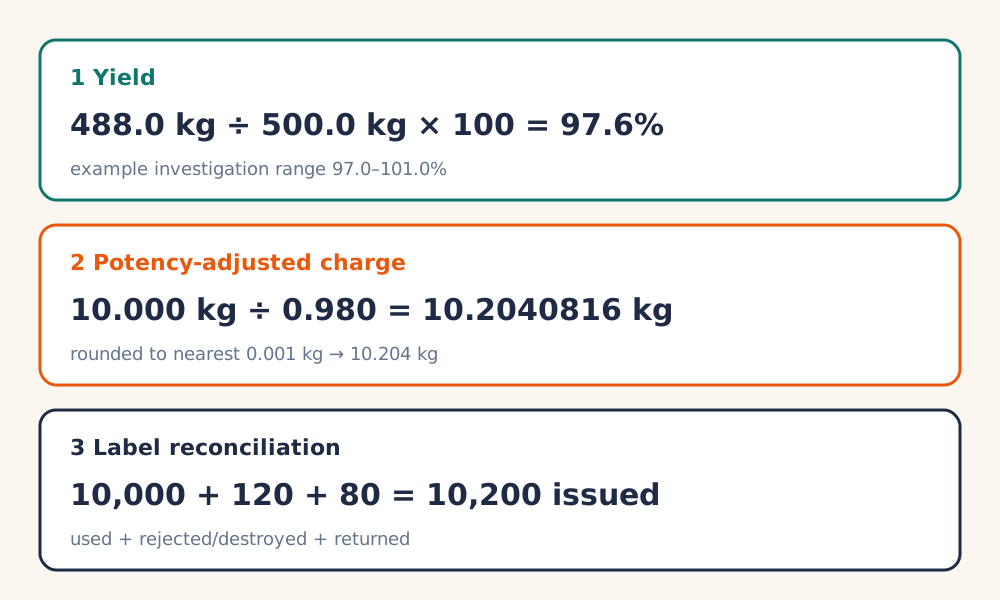
Consider a pharmaceutical process with a theoretical intermediate yield of 500.0 kg and an approved investigation range of 97.0–101.0%. The measured intermediate output is 488.0 kg. The percentage of theoretical yield is 488.0 ÷ 500.0 × 100 = 97.6%. That value falls within this example's approved range. The BMR should preserve the theoretical quantity, actual quantity, calculation and applicable limits rather than recording simply “yield acceptable.”
Now change the measured output to 483.0 kg. The yield becomes 96.6%, below the approved minimum. The record should connect the discrepancy to the required investigation and eventual disposition. A reviewer needs the relevant material, equipment and process context, including potential losses or measurement errors. Do not change the theoretical denominator merely to make the result acceptable. These example limits are assumptions, not universal regulatory thresholds.
A second example concerns a potency-adjusted charge. Suppose an approved formula requires 10.000 kg of active substance on an as-is mass basis. The authorized raw-material assay is 98.0% on that same basis. Under the approved simple compensation method, the required charge is 10.000 ÷ 0.980 = 10.2040816 kg, before applying the process's rounding rule. At a 0.001 kg reporting increment, rounding to nearest gives 10.204 kg.
The BMR should identify the assay result and basis, formula version, calculated target, authorized rounding and measured charge. If the assay is on a dry basis, this simple calculation is not sufficient without the approved basis conversion. If the formula requires a fixed total batch mass, explain how compensation affects other components under the approved formula. A numerically correct active charge can still produce the wrong total composition.
The third example follows labeling quantities. A packaging order issues 10,200 labels. The record accounts for 10,000 used, 120 rejected or destroyed and 80 returned: 10,000 + 120 + 80 = 10,200. The accounting balances under the assumptions of the example. It does not establish that the correct approved artwork was used, that every container received one label or that the returned labels were securely controlled. Assume these are controlled labels without printed lot or control numbers. Under 21 CFR 211.125(d), excess labeling bearing lot or control numbers must be destroyed; it is not stock to return for later use.
Suppose only 60 returned labels are documented. The remaining 20 labels are unexplained. Preserve that discrepancy and its investigation rather than entering a balancing figure. The reviewer also needs approved label identity, packaging quantities and relevant inspections.
Yield reconciliation, strength calculation and labeling accountability should remain distinct controls. One passing calculation cannot substitute for another missing element of batch evidence.
07Data integrity in paper, electronic and hybrid records
The ALCOA+ principles provide a useful framework for assessing whether batch evidence can be trusted. Records should identify their origin, remain understandable, be recorded at the appropriate time, preserve original information and accurately represent the activity. Completeness, consistency, endurance and availability extend that assessment across the record lifecycle. Apply the principles to the evidence itself, not just to the appearance of the final report.
For paper records, controlled issuance, legible entries, attributable corrections and accounting for all pages are important practical controls. A correction should preserve the original entry and explain the change under the applicable procedure. An unexplained rewrite into a clean copy can remove the sequence and circumstances that made the original informative.
For electronic records within Part 11's scope, 21 CFR 11.10 addresses controls for closed systems. These include validation, accurate and complete copies, record protection, authorized access and secure, computer-generated, time-stamped audit trails, among other controls. Part 11 scope is determined through the relevant predicate requirements and use of the record; having a computer in manufacturing does not automatically make every file a Part 11 record.
An audit trail supports understanding of changes. It should not be treated as an alternative to preserving the underlying record. A log saying “value changed” without the relevant value, identity, time and context can leave the reviewer unable to assess the event. Time synchronization and timezone handling also deserve explicit design attention where records cross sites or systems.
E-signatures need a defined meaning and relationship to the signed record. 21 CFR 11.50 addresses signature manifestations, including the signer's name, date and time and meaning of the signature. “Reviewed,” “performed” and “approved” are different assertions. An attractive signature image does not establish what the person approved.
Hybrid records often create the hardest review problem. A paper worksheet may be signed while electronic measurements remain in a scale application and analytical data remain in a laboratory system. Define the authoritative record for each element, how identifiers connect, and which originals and metadata must remain accessible. Data integrity depends on these relationships remaining intact after printing, migration and archiving.
08Deviation management, investigation and release review
A batch record should make exceptions visible without forcing every unusual event into the same severity category. An operator transcription correction, a material identity mismatch, an unapproved substitution and a specification failure raise different questions. Procedures should define how each is recorded, assessed and escalated, while retaining links to the operation and affected material.
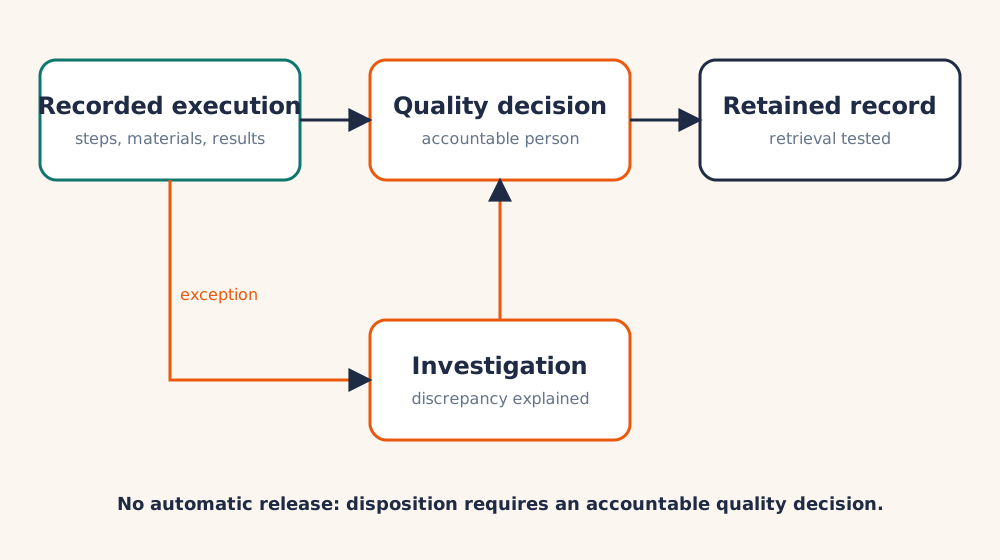
Quality review should establish that the relevant record is complete, applicable instructions were followed or deviations appropriately addressed, results were assessed against the correct criteria and investigations support the proposed disposition. Review is an evidence-based decision, not simply confirmation that every field contains something. A populated field can contain an impossible unit, copied value or result from the wrong sample.
For finished pharmaceuticals, the investigation obligation under 211.192 extends to unexplained discrepancies and specification failures, including potentially associated batches and products. The BMR should therefore support investigation scope: related lots, equipment, production periods and process conditions need to be discoverable. Do not restrict an investigation to the initially selected batch merely because the software's screen shows one record at a time.
A LIMS result and its status should be distinguishable. A preliminary value, authorized result and superseded result can have the same numeric content while different authority. When OOS or OOT matters are investigated, preserve the original data and the justified conclusion. Replacing an initial value with a later passing value can conceal precisely what the reviewer needs to examine.
Batch review by exception can help focus attention when the system reliably identifies relevant exceptions. It needs defined rules, verified data completeness and confidence that missing inputs cannot be classified as clean. Review of exceptions must still support the required quality decision. No rule engine or AI summary acquires the legal authority of the quality unit merely by marking the record green.
Keep investigation closure, CAPA status and batch disposition separate. A long-term CAPA may remain open after an adequately supported batch decision, depending on the circumstances and applicable procedures. Conversely, closing a task does not demonstrate that a batch is suitable for release. Record the rationale and responsible approval for each decision.
Reviewers should be able to move from a summary directly to the supporting evidence. This includes the master version, execution results, source measurements, laboratory records, labeling controls and investigation conclusions.
09ERP, MES, WMS and laboratory integration
A BMR commonly crosses several applications. ERP or material requirements planning software initiates demand and orders; MES coordinates execution; WMS controls material movement; laboratory systems manage samples and tests; QMS supports investigations and quality processes. These are typical architectural roles rather than a requirement that every plant own five separate applications.
Define ownership for each record and status. The component master, approved recipe, lot quality status and authorized analytical result may come from different systems. An integration should communicate the controlled source value, identity, version and relevant timing. Copying fields into another application does not automatically transfer authority over the underlying decision.
Interface failures need a recoverable record. If a consumption transaction is captured in MES but not posted to inventory, the physical consumption has still occurred. Replaying the transaction must avoid double deduction. A stable event identifier, acknowledgment handling and reconciliation process are practical controls for this situation. The batch record should retain the manufacturing event even while the accounting interface is being repaired.
The opposite failure also matters: an ERP order can exist while its approved manufacturing definition fails to load into execution. The presence of an order number must not authorize operators to improvise the missing instructions. Distinguish incomplete interface delivery from process readiness, and define what is required before execution can proceed.
Laboratory integration should preserve the sample-to-result relationship and the approved interpretation. A numeric result alone can omit method, specification version, unit, basis or approval state. Review the meaning of those fields during interface design. A system can successfully transmit “98.0” and still create a dosing error if the receiving formula assumes a different assay basis.
During an outage, record the authorized fallback and subsequent reconciliation. Document which events were captured elsewhere, which were replayed, and how conflicts or duplicate events were resolved. An unexplained gap in machine history cannot be filled by an AI-generated narrative. Where original data are unavailable, the assessment must identify the limitation and its impact.
For implementation, build test cases around meaningful failures as well as normal exchanges: stale master revisions, rejected lots, missing units, delayed results, partial transfers, duplicate messages and corrected source data. Computer system validation should evaluate the intended use and risks of the complete workflow, including interfaces, rather than validating each screen in isolation.
10Designing a usable BMR template and introducing it
Begin with the manufacturing process and review questions, then design the record. Map significant operations, inputs, acceptance criteria, personnel checks and exception paths. Identify where evidence already originates. A process map helps reveal gaps such as a transfer between intermediate containers with no recorded identity or an equipment change that appears in a log but not in the batch history.
A practical template separates batch identity and issuance, material consumption, equipment readiness, processing, sampling and results, packaging, yields, exceptions and disposition. The exact structure should suit the process. A short discrete operation and a multistage pharmaceutical process do not need identical section counts or the same volume of free-text narrative.
For each critical field, define its source, unit, permissible values, required context and review purpose. Consider whether the field captures a target, observation, calculation or decision. This prevents an ambiguous “quantity” field from mixing planned charge, measured weight and booked consumption. It also makes the record more useful for subsequent analysis.
Use structured fields where they improve accuracy, while retaining space for meaningful explanation. A controlled reason code makes recurring causes measurable; a narrative can capture circumstances the code does not describe. Excessive dropdowns can hide complexity just as excessive free text can make analysis difficult. Test the design with operators and reviewers using realistic cases.
Pilot the record on a representative product and workflow. Include an ordinary batch, a changed material lot, a failed measurement, an interruption and a quality exception. Compare the evidence generated with what the reviewer actually needs. A successful demonstration on a perfectly clean batch says little about how the system handles the circumstances that drive investigations.
Training should establish the meaning of entries and signatures, when recording must occur, how errors are corrected and when work must stop. Supervisors and quality reviewers also need to understand the record's limits. A missing alarm history or delayed interface should not be invisible merely because the operator successfully completed the displayed steps.
For an electronic deployment, define intended use, applicable records, configuration ownership, testing and change procedures. Assess data migration and historical access separately from the new workflow. Avoid assuming that an exported PDF is a complete archive of dynamic source data. Implementation is complete when the process produces usable, controlled evidence during both normal work and foreseeable exceptions.
A practical evidence model for the Batch Manufacturing Record
| Evidence | Essential context | Review question |
|---|---|---|
| Material charge | Component and supplier/internal lot identifiers; measured amount, unit, basis and relevant status. | Was the correct eligible material actually used? |
| Process result | Operation, asset, time, value or traceable trend, unit and applicable criteria. | Does the evidence demonstrate the required condition? |
| Laboratory result | Sample identity, method/specification version, result, unit and authorized status. | Does this approved result belong to this batch and sample? |
| Exception | Original observation, scope, investigation reference, conclusion and authorized disposition. | Is the discrepancy explained and its impact assessed? |
| Signature | Identity, time, meaning and the specific record/version signed. | What assertion did this person make? |
11Retention, retrieval and meaningful performance measures
A released batch still needs an accessible manufacturing history. 21 CFR 211.180 establishes record-retention requirements for relevant finished pharmaceutical records, including a rule tied to product expiration and a different provision for certain OTC products without expiration dating. Other product types and jurisdictions have their own requirements. Do not apply a single invented “seven-year GMP rule” to every record.
Create a retention schedule that identifies the record classes, applicable requirements, start events, owners and systems. The BMR summary, source measurements, laboratory records, investigations and relevant metadata may be held in different repositories. Their retention and access arrangements need to support examination of the batch throughout the applicable period.
Backup and archive serve different purposes. A backup helps restore a system after loss; an archive preserves records for later access. Neither is adequate merely because a file exists somewhere. Test retrieval, completeness and readability, including permissions and the relationships between linked evidence. A recovery exercise should also establish whether signatures, audit trails and attachments remain interpretable.
Changes to software or hosting can affect older records. Before retiring a system, identify what is needed to interpret the historical data and how it will remain available. Preserve the relevant versions, context and migration evidence. Repeatedly converting records into new formats without completeness checks can gradually remove information even when each conversion appears visually acceptable.
Measure BMR performance with definitions that support decisions. Review lead time can run from manufacturing completion to quality approval, but record whether laboratory waiting and investigation time are included. Right-first-time records should mean that the record met defined completeness and correctness criteria on first review; simply counting records that reached an approved status can conceal substantial rework.
Track missing entries, corrections, unresolved exceptions, review returns and investigation recurrence. Segment results by product family or operation when process complexity differs. A long processing batch is not directly comparable with a simple packaging order. Avoid incentives that reward closing records quickly while discouraging accurate reporting of unusual events.
Use recurring signals to improve templates, training, interfaces and process controls. A repeated missing tare may indicate poor capture design, while repeated yield discrepancies may point to measurement or process issues. The BMR becomes more valuable when it supports both the individual release decision and a controlled understanding of patterns across manufacturing history.
Frequently asked questions
Q.Is a BMR the same as a master manufacturing record?+
No. The master contains approved instructions and criteria; the BMR contains evidence for a particular batch. They need an explicit version relationship. Changes to the current master should not silently alter what a historical batch appears to have required. Names vary by sector, so procedures should state which document defines the process and which preserves execution.
Q.Does every Batch Manufacturing Record need to be electronic?+
No. Paper records can support compliance when they meet the applicable requirements for content, control, attribution, review and retention. Electronic capture can improve integration and timely checks, but it introduces its own requirements for configuration, validation, access, audit trails and preservation. Select the approach around process risk and intended use rather than assuming digitization itself makes the record reliable.
Q.Can a BMR include links instead of every attachment?+
Often a controlled linked record is appropriate, but the reference must identify the relevant evidence and remain available. Assess whether the reviewer can reconstruct the activity from the linked records and their metadata. A changing URL or file name is insufficient if it can lead to a later version instead of the evidence applicable to the batch.
Q.Who approves a pharmaceutical BMR before release?+
For US finished pharmaceuticals, 21 CFR 211.192 assigns review and approval of production and control records before release or distribution to the quality control unit. The company's workflow can include other operational reviews, but those do not remove the quality unit's responsibility. Other jurisdictions and products may have different release responsibilities, which should be mapped separately.
Q.What happens if the yield is outside the approved range?+
Preserve the measured result and calculation, and initiate the required assessment or investigation under the applicable procedure. For finished pharmaceuticals, the master establishes yield limits that trigger investigation and 211.192 addresses unexplained discrepancies. Determine whether measurement, material, process or other causes are involved. A passing final assay alone does not justify erasing the discrepancy.
Q.Does a PDF contain the whole electronic batch record?+
A PDF may provide an excellent human-readable summary, but completeness depends on what it includes. Source data, audit trails, signature context and dynamic information may remain in other controlled records. Define the complete retained record and test accurate exports and retrieval. Treat printing as a representation of evidence, not automatic proof that every necessary electronic element has been preserved.
Q.Can AI review and release the batch automatically?+
AI can assist with summaries and finding items for attention when its intended use and limitations are controlled. It should not replace accountable quality decisions or invent missing records. Manufacturing discrepancies, analytical results and release authorization require the applicable process and responsible approval. Any AI-produced summary must remain checkable against original evidence rather than becoming an unsupported manufacturing history.
Q.Where should a manufacturer start when records are inconsistent?+
Choose a representative workflow, identify the approved master and map each significant operation to its required evidence. Test whether a reviewer can reconstruct a normal batch and an exception batch. Fix identity, units, version control and source links before adding more fields. Pilot the revised record with operators and quality reviewers, then use the findings to guide a phased rollout.
Primary sources
Further reading
- Master Manufacturing RecordApproved process definition and its relationship to batch execution.
- Electronic Batch Manufacturing RecordElectronic execution, review and record controls.
- Lot GenealogyInput and output relationships across transformations.
- Electronic Batch Record SoftwareV5's electronic batch record product page.
- Batch Record SoftwareV5's batch record product page.
- Sample eBMR batch record (online)Online sample report; opens after the email form.
- Pharmaceutical Industry GuidePharmaceutical operations guide offer; opens after the email form.
Explore this topic
BMR sits inside this topic cluster in our glossary. Every neighbour is one click away.
Master and executed records that prove a batch or device was made to spec.
Want to see how BMR could fit into your own records and workflows? Explore the related V5 pages or talk to our team about what applies to your operation.