Dietary Supplement Packaging and Label Control: Subpart L Operations
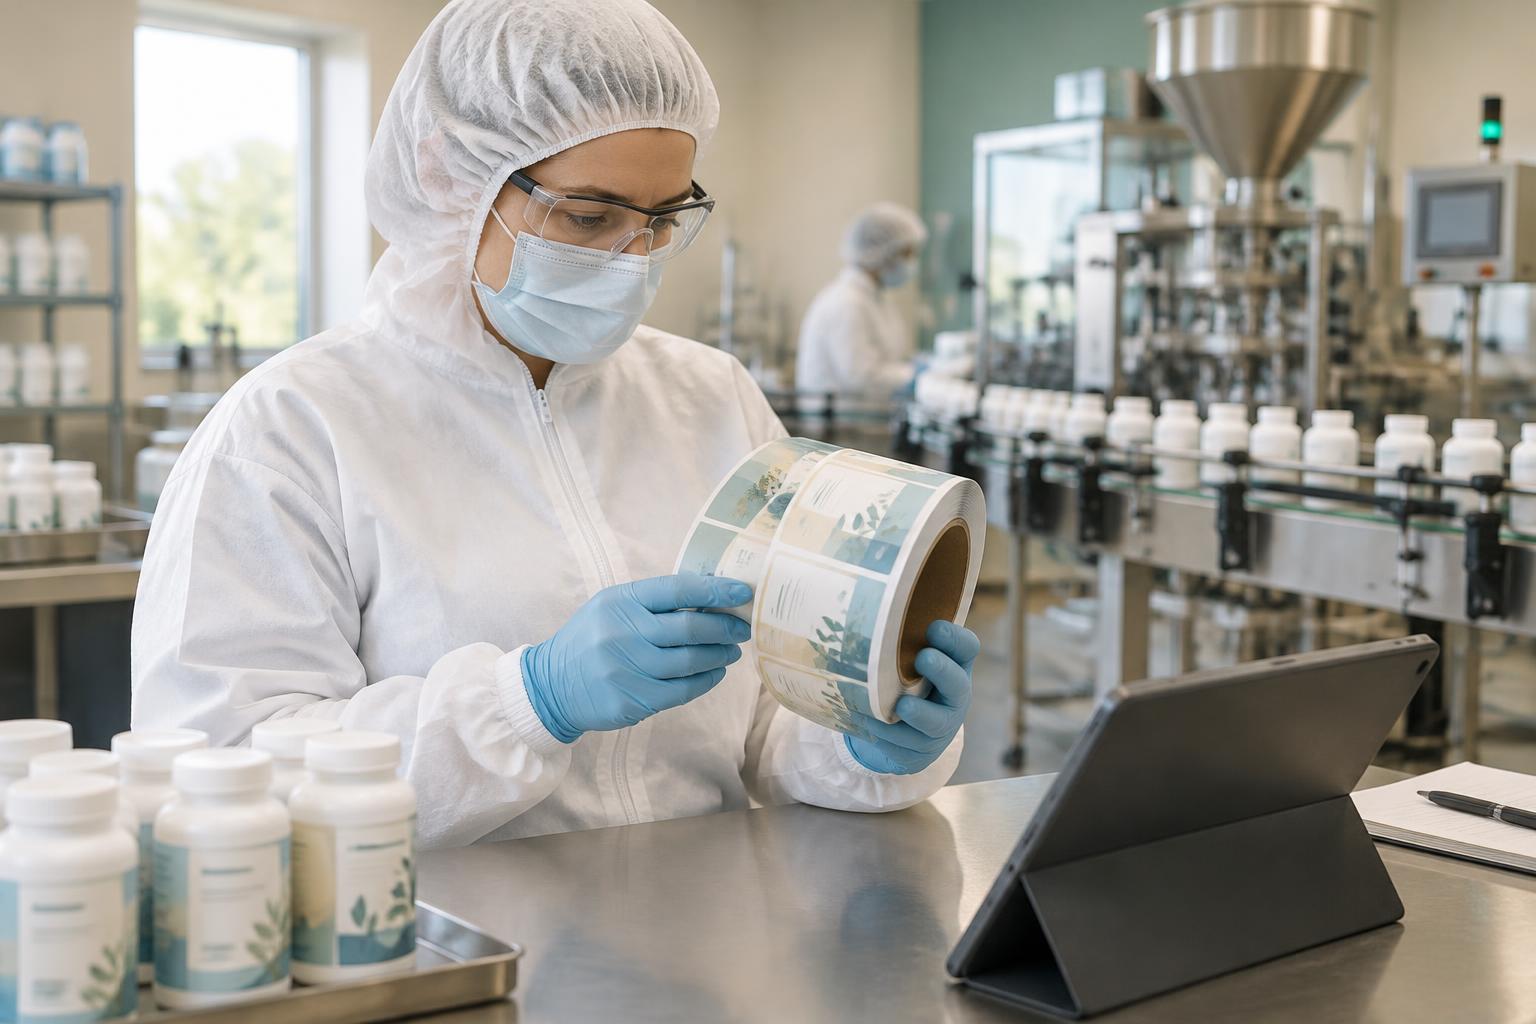
Dietary Supplement Packaging and Label Control connects the packaging line to the approved manufacturing instructions: the right supplement, the specified container and closure, the correct label, and an accountable record of what happened.
In Short
- Dietary Supplement Packaging and Label Control connects the packaging line to the approved manufacturing instructions: the right supplement, the specified container…
- Under §111.70(d), manufacturers must establish label specifications and specifications for packaging that may contact the supplement.
- Written procedures should be usable where the work occurs.
A balanced label count answers one question. Examination against the master manufacturing record answers another. Neither replaces examination of the finished packaged batch or the documented quality control decision.
Control packaging execution without confusing it with label content
Dietary Supplement Packaging and Label Control connects the packaging line to the approved manufacturing instructions: the right supplement, the specified container and closure, the correct label, and an accountable record of what happened. A balanced label count answers one question. Examination against the master manufacturing record answers another. Neither replaces examination of the finished packaged batch or the documented quality control decision.
This guide addresses execution under 21 CFR Part 111 Subpart L, not whether Supplement Facts, ingredient declarations, warnings, or marketing claims satisfy Part 101. It does not interpret those content requirements. For the canonical definition, see packaging and labeling operations under Subpart L; the emphasis here is how operations and evidence fit together.
The starting requirement is straightforward: §111.403 requires written procedures for packaging and labeling operations, and those procedures must be followed. The following map separates the principal operational obligations. References to the CFR identify regulatory requirements; recommendations identified as site practice are implementation choices, not additional universal requirements.
| Section | Operational focus | Evidence to connect |
|---|---|---|
| 111.403 | Establish and follow written procedures | Applicable procedure and execution records |
| 111.410 | Packaging condition; issuance and use; reconciliation; pre-operation MMR conformity; history through distribution | Specifications, issue quantities, examination results, and traceability |
| 111.415 | Filling and packaging practices; separation; identification; sampling; obsolete-material disposal | Handling controls, lot identity, finished-package examination, and disposition |
| 111.420 | QC-approved repackaging or relabeling | Prior approval, representative examination, and QC batch decision |
| 111.425 | Rejected packaged product | Identification, quarantine control, and appropriate disposition |
| 111.430 | Subpart L records | Written procedures and required records maintained under Subpart P |
A useful site practice is to organize the work around five distinct questions: what was authorized, what was issued, what was actually applied, what the finished-package examination found, and what QC decided. Keeping those questions separate prevents a persuasive-looking count sheet from concealing an artwork mismatch, an unidentified container group, or an unresolved packaging defect.
Define the packaging, label, and MMR baseline
Under §111.70(d), manufacturers must establish label specifications and specifications for packaging that may contact the supplement. Contact packaging must be safe and suitable for its intended use and must not be reactive, absorptive, or otherwise affect supplement safety or quality. Section 111.70(g) separately requires finished packaging and labeling specifications, including assurance that the specified packaging and label were used.
Keep three control levels distinguishable: the specification describing acceptable material, the MMR identifying the intended batch configuration, and the batch record documenting execution. The master manufacturing record is the batch-specific comparison baseline for the examination required by §111.410(c). A receiving approval or a familiar product name is not, by itself, that comparison.
| Control level | Question it answers | Site-practice example |
|---|---|---|
| Packaging or label specification | Is this material acceptable for its intended use? | Container material and dimensions; controlled label artwork reference |
| MMR configuration | Is this the configuration specified for this batch? | Container, closure, and label identifiers linked to the applicable MMR revision |
| Finished-package specification | Did the packaged result use the specified materials? | Examination criteria for the completed package and applied label |
| Batch evidence | What was used and what was found? | Material identifiers, quantities, examination results, and QC records |
Before use, §111.75(f) requires minimum examinations and supplier-document review: visual identification of containers and closures, visual examination of labels, and review of the supplier's invoice, guarantee, or certification to determine whether specifications are met. Importantly, §111.75(f) concerns pre-use verification; §111.75(g), not paragraph (f), addresses visual examination of the finished packaged and labeled supplement.
As site practice, assign an unambiguous artwork identifier and revision, and define how an MMR change affects material already received, issued, or staged. Different revision numbers can legitimately coexist across document types: MMR revision 07 need not use label revision 07. The controlled relationship matters. For ingredient-receiving questions, use the separate ingredient identity and supplier CoA guide, rather than treating packaging examination as ingredient identity verification.
Prepare the line with written procedures and separation
Written procedures should be usable where the work occurs. As site practice, specify who retrieves the applicable instructions, who issues materials, how the line is prepared, where examination results are recorded, and who handles exceptions. Define the handoffs between warehouse, packaging, and QC. A procedure that merely says to use the correct label leaves operators to invent the comparison method during a busy changeover.
The operational requirements in §111.415 include cleaning and sanitizing filling and packaging equipment, utensils, and supplement packaging as appropriate; protecting manufactured supplements from contamination, particularly airborne contamination; and sanitary handling. Cleaning controls address contamination. Material-removal and identity checks address mix-ups. Combining them on one form may be convenient, but completing one does not establish that the other occurred.
Section 111.415(d) requires physical or spatial separation of packaging and label operations from operations on other components and supplements to prevent mix-ups. As site practice, translate that into defined staging locations, identifiable work-in-progress areas, and rules for movement between lines. A floor boundary is weak evidence if operators routinely place another batch's labels or open containers inside it.
Line clearance is a practical way to implement mix-up prevention, not a separately named universal checklist in these provisions. A site-designed clearance can examine label unwind and rewind areas, conveyors, coder settings, nearby bins, worktables, and paperwork for remnants of the prior job. Check accessible places where material can remain, rather than treating an empty main work surface as proof that the entire line is ready.
For lot/date-code application, site practice should identify the authorized value, its source, the required format, and how a specimen is compared with instructions. Assigning batch, lot, or control numbers has a specific regulatory basis discussed below; a particular printer setup or date-code workflow is an implementation choice. Do not let a copied setting silently become the authority for the next run.
Control issuance, use, and returns
Section 111.410(b) requires control of packaging and label issuance and use, including reconciliation of issuance-and-use discrepancies. Section 111.410(a) also requires necessary actions to determine whether packaging meets specifications so its condition will ensure supplement quality. Counting labels does not establish that a damaged container or unsuitable closure is acceptable.
As site practice, define an issue transaction with a material identifier, quantity, receiving line or batch, and accountable handoff. Decide how partial rolls, supplemental issues, start-up waste, and returns will be measured. If the warehouse counts rolls while the line counts individual labels, establish the conversion and quantity evidence before the reconciliation is needed. An unexplained unit conversion is not a resolved discrepancy.
- 1. Confirm authorizationIdentify the applicable MMR and specified label.
- MMR identifier and revision
- Label identifier and revision
- 2. Examine before operationsCompare packaging and labels for the batch with the MMR.
- Comparison result
- Material identifiers
- 3. Issue to the lineRecord every issue, including later additions.
- Quantity and unit
- Receiving batch and line
- 4. Record use and lossDistinguish labels applied from labels destroyed.
- Applied quantity
- Documented destruction quantity
- 5. Control remaining materialCount eligible returns and isolate ineligible material for disposition.
- Returned quantity
- Return or disposal status
- 6. Reconcile and resolveCompare issued quantity with accounted quantities; document discrepancies and their resolution.
- Calculated difference
- Supporting evidence and QC review records
Return controls deserve the same care as issuance. As site practice, identify a partial roll before it leaves the line and maintain its revision and quantity status during return. Do not merge an unidentified remainder with available inventory simply because the artwork looks familiar. Material that is obsolete or incorrect belongs in the disposal process, not the reusable-return category.
The regulation contains a narrow exception: label reconciliation is not required for cut or rolled labels if appropriate electronic or electromechanical equipment performs a 100-percent examination for correct labels during or after completion of finishing operations. This is not a universal scanner mandate. It is also not an exception to the surrounding issuance/use controls or to the separate pre-operation examination against the MMR.
Fictional example: reconcile 6,240 labels without overstating the result
The following scenario and quantities are fictional site-practice examples, not prescribed production targets. Packaging lot PK-261018-A runs on line PK-2 under MMR-C60 revision 07, which specifies label LBL-C60 revision 04. The site issues two rolls: LR-A with 3,200 labels and LR-B with 3,040 labels. The procedure uses individual labels as the accounting unit and records all issues to this lot.
For this example, the accounting equation is issued = applied + destroyed + returned. The unexplained difference is issued − applied − destroyed − returned. These categories work because every issued label has exactly one final accounting location; a label cannot be counted as both applied and destroyed.
| Worksheet entry | Quantity | Evidence or calculation |
|---|---|---|
| Roll LR-A issued | 3,200 | Issue record linked to PK-261018-A |
| Roll LR-B issued | 3,040 | Second issue record for the same lot |
| Total issued | 6,240 | 3,200 + 3,040 |
| Applied to containers | 6,012 | Recorded application quantity, including applied labels on held units |
| Destroyed | 37 | 21 start-up labels + 16 labels damaged during running |
| Returned unused | 191 | Counted eligible remainder with label identity retained |
| Total accounted for | 6,240 | 6,012 + 37 + 191 |
| Unexplained difference | 0 | 6,240 − 6,012 − 37 − 191 |
The arithmetic is independently checkable: 6,012 + 37 = 6,049, and 6,049 + 191 = 6,240. Therefore, 6,240 issued = 6,012 applied + 37 destroyed + 191 returned, with zero unexplained difference. Here, applied means a label was attached to a container. It does not mean the container was approved for distribution, and the worksheet does not claim a released-unit total.
Suppose the return record instead showed 181 labels. The difference would be 6,240 − 6,012 − 37 − 181 = 10 labels. As site practice, investigate the issue records, actual remainder, recorded waste, line locations, and counting method. Do not increase the destruction quantity by ten merely to balance the sheet. A corrected record needs evidence for the correction.
Zero reconciliation difference does not prove that the correct artwork or revision was applied. A complete roll of the wrong revision can balance perfectly. The separate §111.410(c) examination against the MMR, performed before packaging and labeling operations, addresses conformity of the packaging and labels. Finished-package examination addresses what is present after execution. Keep both results separate from the quantity balance.
Examine the correct label and the finished packaged batch
Pre-operation examination should answer a precise question: do the packaging and labels for this batch conform to its MMR? As site practice, compare the material identifiers and controlled artwork reference, rather than relying on color, product family, or a label description remembered from a previous run. Where similar presentations exist, identify the specific differences that an examiner needs to distinguish.
The minimum finished-package visual examination appears in §111.75(g): determine whether the specified packaging was used and the specified label applied. Separately, §111.415(g) requires examining a representative sample of each packaged and labeled batch against the specifications established under §111.70(g). These requirements should be connected in the procedure, not confused with the earlier receipt or pre-use checks.
The cited provisions do not supply a universal numerical sample count. Section 111.75(h) requires appropriate, scientifically valid methods for determining whether specifications are met. As site practice, document how the examination approach addresses the batch and relevant opportunities for variation, such as material changes or interrupted operation. Explain the basis for representativeness rather than choosing a convenient count and calling it representative afterward.
Define what the examination records will show. A practical record can identify the packaged lot, applicable specification, units or material examined, observations, result, examiner, and time. An attached image may help communicate an observation, but the site should decide what it actually demonstrates. A photograph of one acceptable bottle does not, by itself, establish how a representative sample was selected or examined.
For sites using the cut/rolled-label reconciliation exception, distinguish examination for the correct label from simple label detection or code readability. As a buyer-evaluation practice, ask how the proposed equipment is assessed for the actual wrong-label risks and how examination coverage is documented. Equipment presence alone does not establish that the conditions of §111.410(b) are met. Nor does electronic examination remove the need to examine packaging and labels against the MMR before operations.
Keep unlabeled containers and packaging lots identifiable
Filled containers held for later labeling create a distinct mix-up risk. Section 111.415(e) requires identification by any effective means when filled containers are set aside and held unlabeled. As site practice, use container-group identification that remains with the material through movement, temporary storage, and division into smaller groups. A nearby note is ineffective if it can become separated from the containers.
Section 111.415(f) requires a batch, lot, or control number for each packaged and labeled lot from a finished batch, and for each lot from a finished batch distributed to another person for packaging or labeling. Keep the finished-batch identity and the resulting packaging-lot identities connected. This is especially important when one bulk batch becomes several package configurations or is handled across separate packaging sessions.
MMR-C60 Revision 07
Specifies → LBL-C60 revision 04 and the authorized packaging configuration.
- Instruction baseline
- Specified material references
Label Rolls LR-a and LR-B
Carry → LBL-C60 revision 04; are issued to → line PK-2 for PK-261018-A.
- Roll identities
- 3,200 and 3,040 issued labels
Packaging Line PK-2
Uses → issued materials; produces → packaged lot PK-261018-A.
- Equipment identity
- Clearance and execution evidence
Packaged Lot PK-261018-A
Derives from → identified finished batch; is examined against → finished-package specifications.
- Finished-batch relationship
- Packaged-lot identity
Batch Production Record
Connects → MMR, rolls, line, packaged lot, examinations, reconciliation, and QC decisions.
- Material and quantity evidence
- Results and disposition records
As site practice, use these relationships to test traceability in both directions. Starting with a packaged lot, can staff identify the applicable instructions, label revision, material identifiers, and examinations? Starting with a suspect roll, can they identify the packaging lots that used it? Separate identifiers are useful only when the records preserve the relationships between them.
Section 111.410(d) requires the ability to determine complete manufacturing history and control through distribution. Linking packaging-lot identifiers to distribution records is a practical implementation approach. Line records that stop at the last carton can establish production activity without, on their own, establishing that broader history.
Handle interruptions, discrepancies, and practical trade-offs
Interruptions can separate identity, quantity, and status. A roll change may introduce another revision; a jam may create waste that never reaches the destruction record; an extended stop may leave unlabeled containers in an ambiguous location. As site practice, identify which events trigger another comparison, a recount, a clearance check, or an assessment of potentially affected material. Do not invent a universal check interval.
The following table describes common failure mechanisms and practical responses. It is not a list of additional CFR clauses. Section 111.75(i) requires corrective action plans for an unmet established specification; the site's response should distinguish a specification failure from a counting or documentation discrepancy while still assessing the implications of either.
| Failure mode | Why a superficial check misses it | Site-practice response |
|---|---|---|
| Wrong revision, balanced count | Reconciliation measures quantity, not artwork conformity | Compare identifiers and artwork with the MMR; assess affected containers |
| Unrecorded supplemental issue | The initial issue total is no longer complete | Capture every addition and recalculate the accountable total |
| Waste counted as applied | A machine count may not represent labels on containers | Define counter meaning and document waste without double counting |
| Unidentified partial roll returned | Product appearance does not establish revision or quantity | Maintain identity and eligibility before returning material to usable inventory |
| Held unlabeled containers moved | Location-based identity may be lost | Keep identification with each moved or divided container group |
| Restart after another job occupied the line | The earlier clearance no longer describes current conditions | Reassess the line and authorized material before resuming |
A practical discrepancy response is to preserve the evidence, identify the potentially affected material, prevent unreviewed disposition under the site's controls, and assess what the discrepancy means. Avoid prematurely describing all exceptions as paperwork errors. A missing label might reflect a count error, unrecorded destruction, or application to an unintended container; those possibilities have different consequences and need different supporting evidence.
Control choices involve trade-offs. Smaller issues simplify the quantity exposed at the line but create more handoffs to record. Dedicated areas reduce shared-space complexity but use more space. Manual examination depends on a clear comparison method; electronic examination adds setup and performance questions. The appropriate choice is the one the site can justify and execute reliably for its actual operation, not the one with the most impressive checklist.
Keep repackaging, rejection, and obsolete-material disposal distinct
Repackaging or relabeling is not permission to quietly restart the original operation. Under §111.420(a), QC personnel must approve repackaging or relabeling before it occurs. As site practice, describe the affected lot and quantity, reason for correction, authorized materials, work instructions, and evidence needed to evaluate the corrected result. Do not perform the work first and seek approval afterward.
Section 111.420(b) requires a representative sample of each repackaged or relabeled batch to be examined against all specifications established under §111.70(g). Section 111.420(c) requires QC to approve or reject each such batch before release for distribution. The approval to undertake corrective work and the decision on the resulting batch are separate decisions; one signature should not be assumed to establish both.
As site practice, preserve the relationship between original execution records and the corrective work. Account for replacement labels and any removed labels without confusing them with the initial run's quantities. If a label is removed from a container, document the transition in a way that preserves the original history and prevents the same label from being counted in incompatible final categories.
A packaged and labeled supplement rejected for distribution has a different status. Section 111.425 requires clear identification, holding, and control under a quarantine system for appropriate disposition. As site practice, ensure the physical location and recorded status agree, and prevent the rejected material from being mistaken for available stock. Quarantine controls the material; it does not itself decide its ultimate disposition.
Obsolete or incorrect labels and packaging must be suitably disposed of so they cannot be used in future packaging and labeling operations, under §111.415(h). As site practice, define the disposal method, responsibility, and evidence appropriate to the material. An obsolete label roll is not an eligible return merely because its quantity is known and its adhesive remains usable.
Assemble batch-record evidence that supports the QC decision
Section 111.255 requires a batch production record each time a supplement batch is manufactured, with complete production and control information, accurate following of the appropriate MMR, and performance of each batch-production step. This guide focuses on the packaging contribution. Use the master and batch records execution guide for the wider record-execution workflow rather than duplicating it in a packaging procedure.
For packaging, §111.260 connects lot numbers, equipment and line identity, cleaning and sanitizing records, and unique material identifiers to batch evidence. Paragraph (k) requires documentation at the time of performance of packaging and labeling operations, including packaging/label identifiers and quantities, label-discrepancy reconciliation when required, an actual or representative label or its specified location, and examination results or their physical location.
As site practice, arrange those records so a reviewer can follow the event sequence without reconstructing it from disconnected totals. Cross-references should identify the correct record and location, not merely say see log. Keep the approved label reference distinguishable from the actual or representative label evidence. One expresses the intended configuration; the other helps document the batch's packaging and labeling record.
Section 111.105 establishes QC responsibilities for quality and conformity to the MMR, including specified approval/rejection activities and ensuring required representative samples are collected. For laboratory operations associated with the production and process control system, §111.110 requires QC to ensure the §111.75 tests and examinations are conducted and to review and approve their results. Do not reduce QC review to checking arithmetic alone.
Section 111.260(l) requires contemporaneous documentation of QC batch-record review and the relevant approval, release, or rejection decisions, including for packaged and labeled supplements. Section 111.260(m) covers any required material-review and disposition decision. Finally, §111.430 requires Subpart L records, including written packaging and labeling procedures, to be made and kept in accordance with Subpart P. This guide does not restate Subpart P's retention rules.
Implement a finite packaging evidence checklist
Start with one defined packaging configuration rather than attempting to redesign every record simultaneously. As site practice, walk its instructions from material availability to the QC decision and distribution linkage. Identify where the site currently proves identity, quantity, condition, examination, and status. Then address specific missing connections. Adding a second signature to an unclear record rarely fixes an undefined quantity or an ambiguous material identifier.
The following ten-item checklist is a practical evidence review, not a replacement for the regulations or the site's approved procedures. It deliberately separates mandatory outcomes from the local forms used to demonstrate them. Apply it to a completed packaging job and, where relevant, to an interrupted or corrected job so that the review tests more than an uncomplicated run.
- Procedure baseline: identify the applicable written packaging and labeling procedures and where execution is documented.
- Specification baseline: connect contact-packaging and label specifications with finished-package specifications and the applicable MMR.
- Pre-operation examination: locate the batch-specific packaging and label comparison against the MMR, separately from receiving checks.
- Line readiness: locate site-practice clearance evidence and the applicable cleaning, handling, and separation controls.
- Issue accountability: account for initial and supplemental issues using defined quantities, units, and material identifiers.
- Identity continuity: trace unlabeled container groups, the finished batch, and each resulting packaged lot.
- Quantity conclusion: locate reconciliation and discrepancy resolution, or establish the basis for the narrow label-reconciliation exception.
- Finished-package conclusion: locate representative-sample examination results against the established packaging and labeling specifications.
- Exception disposition: locate applicable repackaging/relabeling approvals, corrected-batch examinations, rejection controls, and obsolete-material disposition.
- QC and record closure: locate the documented QC decisions, required batch evidence, retrievable cross-references, and distribution linkage.
Use gaps to improve the underlying control, not merely the wording of the checklist. An uncertain return quantity needs a better return-counting method; an ambiguous revision needs a clearer authorization relationship. The practical objective is a coherent account of what was intended, used, examined, and decided. A neat reconciliation sheet is valuable evidence, but it is only one part of that account.
Questions
Does Subpart L require a scanner on every packaging line?
No universal scanner requirement appears in §111.410(b). It provides a narrow label-reconciliation exception for cut or rolled labels when appropriate electronic or electromechanical equipment performs a 100-percent examination for correct labels during or after finishing operations.
Does zero reconciliation difference prove the correct label was used?
No. It establishes a quantity balance under the accounting method used. The separate §111.410(c) pre-operation examination determines packaging and label conformity to the MMR; finished-package examination addresses the packaged result.
Is finished-package visual examination in §111.75(f)?
No. Paragraph (f) addresses pre-use packaging and label examinations and supplier-document review. Paragraph (g) requires the minimum visual examination of finished packaging and labeling. Section 111.415(g) also requires representative-sample examination of each packaged and labeled batch.
What sample size must be used for packaged-batch examination?
The cited provisions do not prescribe a universal numerical count. They require representative-sample examination and appropriate, scientifically valid methods. As site practice, document the selection approach and why it is suitable for the batch and examination.
Is line clearance an explicitly named Subpart L requirement?
These provisions require effective packaging practices and mix-up prevention, including physical or spatial separation. A documented line-clearance workflow is a site practice for implementing those controls, not a universally prescribed checklist in the cited text.
Can operators relabel a batch before QC approval?
Section 111.420 permits relabeling or repackaging only after QC approval. It also requires representative-sample examination of the corrected batch and QC approval or rejection before release for distribution.
Does this guide explain Supplement Facts or marketing-claim compliance?
No. It addresses packaging and label execution under Part 111, not Part 101 content interpretation. The operational question is whether the specified label was controlled and applied; the legality of its content requires a separate assessment.
Sources
- 21 CFR 111.403 — Written packaging and labeling procedures
- 21 CFR 111.410 — Packaging and label requirements
- 21 CFR 111.415 — Filling, packaging, labeling, and related operations
- 21 CFR 111.420 — Repackaging and relabeling
- 21 CFR 111.425 — Rejected packaged and labeled supplements
- 21 CFR 111.430 — Subpart L records
- 21 CFR 111.70 — Specifications
- 21 CFR 111.75 — Determining whether specifications are met
- 21 CFR 111.105 — Quality control responsibilities
- 21 CFR 111.110 — QC laboratory operations
- 21 CFR 111.255 — Batch production record requirements
- 21 CFR 111.260 — Batch production record contents
Practical orientation, not legal advice. Rules differ by jurisdiction; your quality and compliance team decides what applies.
The Dietary Supplement Operations Guide
What the label says. What the batch proves.
- The rules and who decides
- Specification to finished batch
- Test, review, investigate
- Labels, samples and recall
Plus Evidence map and four worksheets.
Get the supplement guideWe ask for your name and email so we can handle this request. The full guide then opens on this site, ready to translate, print or save. Privacy policy. Practical orientation, not legal advice; your quality unit decides which rules apply.
 Ultimate
Free guide
The Dietary Supplement Operations Guide
What the label says. What the batch proves.
Keep reading
 Dietary Supplement cGMP Readiness: Handoffs and Audit Evidence
 21 CFR 111 Readiness: Identity, Master and Batch Records
 NDI Notification under 21 CFR 190.6: Contents, the 75-Day Wait and Amendments
 NSF 173 & USP <2750> Readiness: Third-Party Supplement Certification
 Structure/Function Claims Readiness: DSHEA §6, Disclaimers & Substantiation
 Supplement Complaint and Adverse Event Reporting Readiness: U.S. Guide