EU GMP Annex 1: show how every sterile control fits together
Annex 1 asks one thing above all: can you show why each contamination control exists, and that it works?
In Short
- The contamination control strategy links every control to a risk.
- Aseptic process simulation: normally twice a year per line and shift.
- The qualified person decides certification, not the monitoring system.
The Pharmaceutical Operations Guide
Right material. Right record. Released by a person.
- Who decides, and who may not
- Supplier to finished bulk
- Test, review, investigate
- Packs, serialisation and recall
Plus Evidence map and four worksheets.
Get the pharmaceutical guideWe ask for your name and email so we can handle this request. The full guide then opens on this site, ready to translate, print or save. Privacy policy. Practical orientation, not legal advice; your quality unit decides which rules apply.
 Ultimate
Free guide
The Pharmaceutical Operations Guide
Right material. Right record. Released by a person.
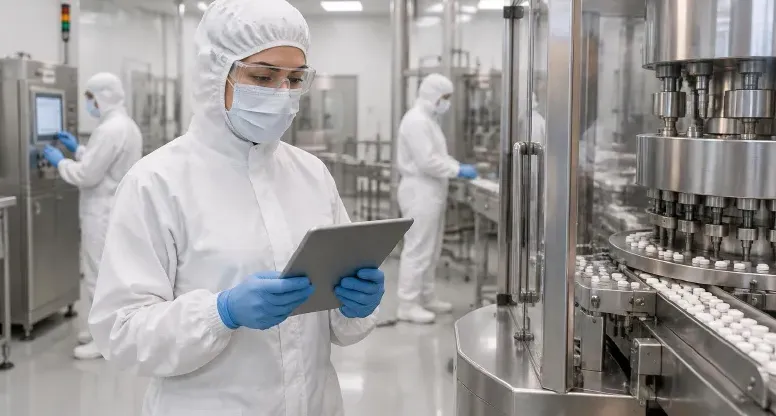
Annex 1 of the EU GMP Guide covers the manufacture of sterile medicinal products. The 2022 revision is listed in EudraLex Volume 4 as fully applicable since 25 Aug 2024. It is Commission guidance that inspectors use to judge GMP compliance; PIC/S authorities adopted the same text. This guide covers what sterile QA, microbiology and engineering teams ask about most. Paragraph numbers were checked against the official Annex 1 PDF on 2 Oct 2026.
The Contamination Control Strategy
Annex 1 asks for a contamination control strategy (CCS) across the facility. Paragraph 2.5 lists what it should consider, including plant and process design, premises and equipment, personnel, utilities, raw materials, product containers, vendor approval, process risk assessment, process validation, preventive maintenance, cleaning and disinfection, monitoring systems, root cause investigation and CAPA, and continuous improvement.
Trace one control back to its risk
Pick a control to see the contamination risk your strategy should tie it to.
- Gowning qualification, glove checks and personnel monitoring results.
More on the Contamination Control Strategy
The point is that each control is justified by a contamination risk, and that the strategy is reviewed and kept current.
Cleanroom Grades and Monitoring
Grades A to D set particle and microbial limits at rest and in operation. Grade A is the zone for high-risk operations such as aseptic filling.
More on cleanroom Grades and Monitoring
Monitoring locations, frequency and alert and action limits should come from risk assessment and qualification data. Results trend over time; a single excursion is investigated, and a run of alerts is treated as a signal even when no action limit is breached.
Aseptic Process Simulation
Paragraph 9.38 says APS should be part of initial validation with at least three consecutive satisfactory runs covering all working shifts, then normally repeated twice a year (about every six months) for each aseptic process, filling line and shift. Each operator should take part in at least one successful APS annually.
Aseptic process simulation cadence (9.38)
| When | What Annex 1 expects |
|---|---|
| Initial validation | At least three consecutive satisfactory runs covering all shifts |
| Routine | Normally twice a year (about every six months) per process, line and shift |
| Each operator | At least one successful APS annually |
| After significant change | Repeat where the change affects sterility assurance, e.g. HVAC or shift pattern |
- Initial validation
- What Annex 1 expects
- At least three consecutive satisfactory runs covering all shifts
- Routine
- What Annex 1 expects
- Normally twice a year (about every six months) per process, line and shift
- Each operator
- What Annex 1 expects
- At least one successful APS annually
- After significant change
- What Annex 1 expects
- Repeat where the change affects sterility assurance, e.g. HVAC or shift pattern
More on aseptic Process Simulation
The simulation should imitate the routine process as closely as possible, including interventions.
Sterilising Filters and PUPSIT
Paragraph 8.87 expects the integrity of the sterilised filter assembly to be verified before use (pre-use post-sterilisation integrity test, PUPSIT) and a non-destructive integrity test after use. The annex recognises that PUPSIT may not always be possible after sterilisation because of process constraints, such as filtering very small volumes; an alternative approach may then be taken after a thorough risk assessment.
Records and who decides
Annex 1 relies on records: environmental and personnel monitoring, APS results, filter tests, sterilisation cycles, interventions and deviations. The qualified person certifies batches.
Worked Example: a Grade a Excursion
- 10:42Settle plate at filling needle zone shows growth after incubationMicrobiology
- Day 0Deviation DV-2214 opened; batch S-0917 stays on holdQA
- Day 2Organism identified; interventions and gowning records for the shift reviewedMicrobiology, QA
- Day 6Cause: unrecorded intervention; CAPA raised for intervention trainingQA
- Day 7Qualified person records the batch decision and rationaleQualified person
More on records and who decides
Software can collect and link the evidence, but people decide whether an excursion affects a batch and whether it may be certified.
Questions
Is Annex 1 law?
It is guidance within the EU GMP Guide (EudraLex Volume 4). EU law requires manufacturers to follow GMP, and inspectors use the annex to judge whether they do. The 2022 revision has been fully applicable since 25 Aug 2024.
How often should aseptic process simulation be repeated?
Paragraph 9.38 says normally twice a year, about every six months, for each aseptic process, filling line and shift, after initial validation of at least three consecutive satisfactory runs. Each operator should take part in at least one successful APS each year.
Is PUPSIT always required?
Annex 1 expects it. Paragraph 8.87 recognises it may not always be possible because of process constraints, and allows an alternative approach after a thorough risk assessment.
Does V5 decide whether an excursion affects a batch?
No. V5 keeps deviations, CAPA, batch records and controlled documents with electronic signatures. Your microbiology and QA teams investigate, and the qualified person decides certification.
Practical orientation, not legal advice. Rules differ by jurisdiction; your quality and compliance team decides what applies.
Keep reading
 EU GMP Annex 11 Computerised Systems Guide (2011 text and 2025 draft)
 EU AI Act (Regulation 2024/1689) for Regulated Manufacturers
 EU GMP for Medicinal Cannabis: GACP, Annex 7 and Herbal Starting Materials
 EU GMP Guide: EudraLex Volume 4 Chapters, Annexes and Who Decides
 GMP Manufacturing Readiness: a Practical Batch Evidence Walk
 EU Acrylamide Regulation 2017/2158: Mitigation, Sampling and Benchmark Levels